
D-Threose
Athanor · official
C4H8O4 · 120.1 g/mol
IUPAC name: (2R,3S)-2,3,4-trihydroxybutanal
Also called tetrose
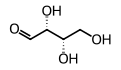
Names and identifiers
- IUPAC name
- (2R,3S)-2,3,4-trihydroxybutanal
- Synonyms
- tetrose, L-Threose, Threose, L-
+ 30 more
L-Threofuranose, MHH79K1BVR, CHEBI:21405, DTXSID40915184, RefChem:397532, GlyTouCan:G77428YW, DTXCID901475619, G77428YW, 202-419-6, (2R,3S)-2,3,4-trihydroxybutanal, L-threo-tetrose, UNII-MHH79K1BVR, (2R,3S)-2,3,4-Trihydroxy-butanal; [R-(R*,S*)]-2,3,4-Trihydroxy-butanal; L-Threofuranose, EINECS 202-419-6, L-(+)-Threosesyrup, L-THREOSE [MI], L-Threose - aqueous solution, SCHEMBL117909, orb2939643, SCHEMBL7718568, YTBSYETUWUMLBZ-IMJSIDKUSA-N, DTXSID601017426, GEO-04659, TYD-00954, AKOS016844121, CS-W021028, HY-W040288, MT06722, Q27109399, F49E7756-40E6-4325-AABD-CAE8087B66F9
- Molecular formula
- C4H8O4
- Empirical formula
- CH2O
- Molecular weight
- 120.1 g/mol
- Exact mass
- 120.04226
- CAS number
- 95-43-2
- PubChem CID
- 5460672
- Canonical SMILES
- C(C(C(C=O)O)O)O
- Isomeric SMILES
- C([C@@H]([C@H](C=O)O)O)O
- InChI
- InChI=1S/C4H8O4/c5-1-3(7)4(8)2-6/h1,3-4,6-8H,2H2/t3-,4-/m0/s1
- InChIKey
- YTBSYETUWUMLBZ-IMJSIDKUSA-N
- Molfile
Names, InChI and the identifiers from PubChem, retrieved 2026-10-14.
Calculated
- Heavy atoms
- 8
- Rings
- 0
- Aromatic rings
- 0
- Fraction sp³ carbons (Fsp3)
- 0.75
- H-bond donors
- 3
- H-bond acceptors
- 4
- Rotatable bonds
- 3
- TPSA (Ų)
- 77.8
- Formal charge
- 0
- XLogP3
- -2.2 PubChem
- Complexity
- 72 PubChem
- Stereocenters
- 2 definedCIP: C3: S · C5: R
- Double-bond geometry
- None specified
- Point group
- C1 of one 3D conformer with its hydrogens, by Athanor’s engine (as in the app)
Elemental analysis
- C40.00%
- H6.71%
- O53.28%
Isotope pattern
Each peak is a nominal mass with its abundance-weighted exact mass, as a percentage of the tallest, from IUPAC natural isotope abundances.
Computed from the structure with RDKit 2026.09 when this page was built, beside PubChem's own computed values from its record; where the two differ, both are shown.
Predicted
- log P
- -2.1 Crippen (Wildman and Crippen 1999)
- log S (water solubility)
- 0.94 ESOL (Delaney 2004), log mol/L
- Synthetic accessibility
- 3.8 SA score, 1 easy to 10 hard (Ertl and Schuffenhauer 2009)
Drug-likeness: Lipinski’s rule of five
- Molecular weight ≤ 500: 120.1
- log P ≤ 5 (Crippen): -2.1
- H-bond donors ≤ 5: 3
- H-bond acceptors ≤ 10: 4
Passes all four rules.
¹³C NMR shifts, predicted
Predictions, not measurements; each names its method. A measured log P, where a source reports one, is under Physical properties.
Safety
No GHS hazard classification was found in the sources.
Safety data sheets on the supplier’s page: AKos Consulting & Solutions, AA BLOCKS, EvitaChem.
Always read the supplier’s safety data sheet before working with it.
Literature
Reading PubChem, OpenAlex and GSRS…
Bioactivity
Reading ChEMBL…
References
- ·Calculated values: computed by Athanor with RDKit 2026.09 from the structure.
- ·Literature on PubChem
Data last updated 2026-10-14.