
D-Lysine
Athanor · official
C6H14N2O2 · 146.19 g/mol
IUPAC name: (2R)-2,6-diaminohexanoic acid
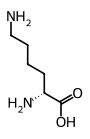
Names and identifiers
- IUPAC name
- (2R)-2,6-diaminohexanoic acid
- Synonyms
- (R)-2,6-Diaminohexanoic acid, (2R)-2,6-diaminohexanoic acid, D-2,6-Diaminohexanoic acid
+ 34 more
Lysine, D-, D-Lysin, (r)-lysine, UNII-3HQ6U6424Q, 3HQ6U6424Q, D-2,6-Diaminohexanoate, EINECS 213-091-9, (-)-R-LYSINE, DTXSID4074986, CHEBI:16855, RefChem:130646, (R)-2,6-Diaminohexanoate, DTXCID8048789, 213-091-9, KDXKERNSBIXSRK-RXMQYKEDSA-N, MFCD00008234, D-Lys, DLY, Lysine #, HSDB 2108, Aminutrin, 6-amino-, L-Lysine (9CI), D-Lysine(>90%), Lysine, L- (8CI), SCHEMBL7343, CHEMBL319497, GTPL4681, orb2944196, SCHEMBL29438737, BDBM217368, D-Lysine, >=98% (HPLC), (R,S)-2,6-Diamino-hexanoic acid, HY-Y1091, EBC-44077
- Molecular formula
- C6H14N2O2
- Empirical formula
- C3H7NO
- Molecular weight
- 146.19 g/mol
- Exact mass
- 146.10553
- CAS number
- 923-27-3
- PubChem CID
- 57449
- Canonical SMILES
- C(CCN)CC(C(=O)O)N
- Isomeric SMILES
- C(CCN)C[C@H](C(=O)O)N
- InChI
- InChI=1S/C6H14N2O2/c7-4-2-1-3-5(8)6(9)10/h5H,1-4,7-8H2,(H,9,10)/t5-/m1/s1
- InChIKey
- KDXKERNSBIXSRK-RXMQYKEDSA-N
- Molfile
Names, InChI and the identifiers from PubChem, retrieved 2026-10-14.
Calculated
- Heavy atoms
- 10
- Rings
- 0
- Aromatic rings
- 0
- Fraction sp³ carbons (Fsp3)
- 0.83
- H-bond donors
- 3
- H-bond acceptors
- 3 computed here · PubChem 4
- Rotatable bonds
- 5
- TPSA (Ų)
- 89.3
- Formal charge
- 0
- XLogP3
- -3 PubChem
- Complexity
- 106 PubChem
- Stereocenters
- 1 definedCIP: C6: R
- Double-bond geometry
- None specified
- Point group
- C1 of one 3D conformer with its hydrogens, by Athanor’s engine (as in the app)
Elemental analysis
- C49.30%
- H9.65%
- N19.16%
- O21.89%
Isotope pattern
Each peak is a nominal mass with its abundance-weighted exact mass, as a percentage of the tallest, from IUPAC natural isotope abundances.
Computed from the structure with RDKit 2026.09 when this page was built, beside PubChem's own computed values from its record; where the two differ, both are shown.
Predicted
- log P
- -0.47 Crippen (Wildman and Crippen 1999)
- log S (water solubility)
- -0.12 ESOL (Delaney 2004), log mol/L
- Synthetic accessibility
- 2.47 SA score, 1 easy to 10 hard (Ertl and Schuffenhauer 2009)
Drug-likeness: Lipinski’s rule of five
- Molecular weight ≤ 500: 146.19
- log P ≤ 5 (Crippen): -0.47
- H-bond donors ≤ 5: 3
- H-bond acceptors ≤ 10: 3
Passes all four rules.
¹³C NMR shifts, predicted
Predictions, not measurements; each names its method. A measured log P, where a source reports one, is under Physical properties.
Safety
This chemical does not meet GHS hazard criteria for 100% (2 of 2) of all reports.[1]
Safety data sheets on the supplier’s page: Oakwood Products, Aladdin Scientific, BOC Sciences.
Always read the supplier’s safety data sheet before working with it.
Spectra
- NMRNMRShiftDB
- 1H NMRHuman Metabolome Database (HMDB) (Bruker · 600 MHz · Water)
- 13C NMRSpectraBase (Bruker WP-80)
- 1H-13C NMRHuman Metabolome Database (HMDB) (Bruker · 600 MHz · Water)
- GC-MSMassBank of North America (MoNA) (GC-MS) · MassBank of North America (MoNA) (GC-MS) · MassBank of North America (MoNA) (GC-MS) · NIST Mass Spectrometry Data Center
- MS-MSHuman Metabolome Database (HMDB) (LC-ESI-ITFT (LTQ Orbitrap XL, Thermo Scientfic) · Positive) · Human Metabolome Database (HMDB) (LC-ESI-ITFT (LTQ Orbitrap XL, Thermo Scientfic) · Positive) · Human Metabolome Database (HMDB) (LC-ESI-ITFT (LTQ Orbitrap XL, Thermo Scientfic) · Positive) · Human Metabolome Database (HMDB) (LC-ESI-ITFT (LTQ Orbitrap XL, Thermo Scientfic) · Positive) · Human Metabolome Database (HMDB) (LC-ESI-ITFT (LTQ Orbitrap XL, Thermo Scientfic) · Positive) · Human Metabolome Database (HMDB) (LC-ESI-ITFT (LTQ Orbitrap XL, Thermo Scientfic) · Positive)
- LC-MSMassBank of North America (MoNA) (Quattro_QQQ · positive) · MassBank of North America (MoNA) (Quattro_QQQ · positive) · MassBank of North America (MoNA) (Quattro_QQQ · positive)
- ATR-IRSpectraBase
Experimental spectra held by these sources; each link opens the source's own record.
Literature
Reading PubChem, OpenAlex and GSRS…
Bioactivity
Reading ChEMBL…
References
- [1]European Chemicals Agency (ECHA) · ECHA legal notice: reuse with acknowledgement · via PubChem · retrieved 2026-10-14
- [2]NMRShiftDB · Linked only · via PubChem · retrieved 2026-10-14
- [3]Human Metabolome Database (HMDB) · Linked only · via PubChem · retrieved 2026-10-14
- [4]SpectraBase · Linked only · via PubChem · retrieved 2026-10-14
- [5]MassBank of North America (MoNA) · CC BY 4.0 · via PubChem · retrieved 2026-10-14
- [6]NIST Mass Spectrometry Data Center · Linked only · via PubChem · retrieved 2026-10-14
- ·Calculated values: computed by Athanor with RDKit 2026.09 from the structure.
- ·Literature on PubChem
Data last updated 2026-10-14.