
GABA
Athanor · official
C4H9NO2 · 103.12 g/mol
IUPAC name: 4-aminobutanoic acid
Also called gamma-aminobutyric acid, 4-aminobutanoic acid, piperidinic acid
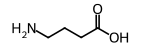
Names and identifiers
- IUPAC name
- 4-aminobutanoic acid
- Synonyms
- gamma-aminobutyric acid, 4-aminobutanoic acid, piperidinic acid
+ 37 more
Piperidic acid, Piperidinic acid, Aminalon, Gaballon, Mielomade, Gamarex, Gammalon, Gammar, Gammasol, butanoic acid, 4-amino-, Gammalone, Butyric acid, 4-amino-, gamma-Aminobutanoic acid, Immu-G, gamma-amino-n-butyric acid, Gammagee, Gamulin, 4-aminobutyrate, gamma-Aminobuttersaeure, omega-Aminobutyric acid, 4-aminobutanoate, Aminalone, Factor I, 4Abu, gamma Aminobutyric acid, 2ACZ6IPC6I, Aminobutyric acid, gamma-, NSC-27418, CHEBI:16865, Acid, Aminobutyric, gamma-amino-butyric acid, Therafeldamine, Theraproxen, GABA Phenolic, Therapentin-60, Theraprofen-60, Theraprofen-90
- Molecular formula
- C4H9NO2
- Empirical formula
- C4H9NO2
- Molecular weight
- 103.12 g/mol
- Exact mass
- 103.06333
- CAS number
- 56-12-2
- PubChem CID
- 119
- Canonical SMILES
- C(CC(=O)O)CN
- Isomeric SMILES
- C(CC(=O)O)CN
- InChI
- InChI=1S/C4H9NO2/c5-3-1-2-4(6)7/h1-3,5H2,(H,6,7)
- InChIKey
- BTCSSZJGUNDROE-UHFFFAOYSA-N
- Molfile
Names, InChI and the identifiers from PubChem, retrieved 2026-10-14.
Calculated
- Heavy atoms
- 7
- Rings
- 0
- Aromatic rings
- 0
- Fraction sp³ carbons (Fsp3)
- 0.75
- H-bond donors
- 2
- H-bond acceptors
- 2 computed here · PubChem 3
- Rotatable bonds
- 3
- TPSA (Ų)
- 63.3
- Formal charge
- 0
- XLogP3
- -3.2 PubChem
- Complexity
- 62 PubChem
- Stereocenters
- None
- Double-bond geometry
- None specified
- Point group
- C1 of one 3D conformer with its hydrogens, by Athanor’s engine (as in the app)
Elemental analysis
- C46.59%
- H8.80%
- N13.58%
- O31.03%
Isotope pattern
Each peak is a nominal mass with its abundance-weighted exact mass, as a percentage of the tallest, from IUPAC natural isotope abundances.
Computed from the structure with RDKit 2026.09 when this page was built, beside PubChem's own computed values from its record; where the two differ, both are shown.
Predicted
- log P
- -0.19 Crippen (Wildman and Crippen 1999)
- log S (water solubility)
- -0.16 ESOL (Delaney 2004), log mol/L
- Synthetic accessibility
- 1.85 SA score, 1 easy to 10 hard (Ertl and Schuffenhauer 2009)
Drug-likeness: Lipinski’s rule of five
- Molecular weight ≤ 500: 103.12
- log P ≤ 5 (Crippen): -0.19
- H-bond donors ≤ 5: 2
- H-bond acceptors ≤ 10: 2
Passes all four rules.
¹³C NMR shifts, predicted
Predictions, not measurements; each names its method. A measured log P, where a source reports one, is under Physical properties.
Safety
No GHS hazard classification was found in the sources.
Safety data sheets on the supplier’s page: SynHet - Synthetic Heterocycles, Aladdin Scientific, Alfa Chemistry.
Always read the supplier’s safety data sheet before working with it.
Spectra
- NMRNMRShiftDB
- 1H NMRHuman Metabolome Database (HMDB) (Varian · 600 MHz · Water) · Human Metabolome Database (HMDB) (JEOL · 90 MHz · D2O) · Human Metabolome Database (HMDB) (Bruker · D2O) · SpectraBase
- 13C NMRHuman Metabolome Database (HMDB) (Bruker · 125 MHz · Water) · Human Metabolome Database (HMDB) (400 MHz · H2O) · Human Metabolome Database (HMDB) (Varian · 25.16 MHz · D2O) · SpectraBase
- 1H-13C NMRHuman Metabolome Database (HMDB) (Bruker · 600 MHz · Water)
- GC-MSHuman Metabolome Database (HMDB) (GC-MS) · Human Metabolome Database (HMDB) (GC-MS) · Human Metabolome Database (HMDB) (EI-B · positive) · Human Metabolome Database (HMDB) (EI-B · positive) · Human Metabolome Database (HMDB) (GC-EI-TOF · positive) · Human Metabolome Database (HMDB) (GC-EI-TOF · positive)
- MS-MSHuman Metabolome Database (HMDB) (Quattro_QQQ · Positive) · Human Metabolome Database (HMDB) (Quattro_QQQ · Positive) · Human Metabolome Database (HMDB) (Quattro_QQQ · Positive) · Human Metabolome Database (HMDB) (Positive) · Human Metabolome Database (HMDB) (Positive) · Human Metabolome Database (HMDB) (Positive)
- LC-MSMassBank Europe (API3000, Applied Biosystems · LC-ESI-QQ · NEGATIVE) · MassBank Europe (API3000, Applied Biosystems · LC-ESI-QQ · NEGATIVE) · MassBank Europe (API3000, Applied Biosystems · LC-ESI-QQ · NEGATIVE) · MassBank Europe (API3000, Applied Biosystems · LC-ESI-QQ · POSITIVE) · MassBank Europe (API3000, Applied Biosystems · LC-ESI-QQ · POSITIVE) · MassBank Europe (API3000, Applied Biosystems · LC-ESI-QQ · POSITIVE)
- Other MSMassBank Europe (HITACHI M-2500 · EI-B · POSITIVE) · MassBank Europe (GCMS-2010 Plus, Shimadzu · EI-B · POSITIVE) · MassBank Europe (waters xevo g2 TOF · ESI-TOF · POSITIVE) · MassBank Europe (waters xevo g2 TOF · ESI-TOF · POSITIVE) · MassBank of North America (MoNA) (TQD, Waters · Flow-injection QqQ/MS · ESI) · MassBank of North America (MoNA) (TQD, Waters · Flow-injection QqQ/MS · ESI)
- FTIRSpectraBase (Bruker IFS 85 · KBr-Pellet)
- ATR-IRSpectraBase (Bio-Rad FTS · ATR-Neat (DuraSamplIR II))
- RamanSpectraBase (FT-Raman)
Experimental spectra held by these sources; each link opens the source's own record.
Literature
Reading PubChem, OpenAlex and GSRS…
Bioactivity
Reading ChEMBL…
References
- [1]NMRShiftDB · Linked only · via PubChem · retrieved 2026-10-14
- [2]Human Metabolome Database (HMDB) · Linked only · via PubChem · retrieved 2026-10-14
- [3]SpectraBase · Linked only · via PubChem · retrieved 2026-10-14
- [4]MassBank Europe · Linked only · via PubChem · retrieved 2026-10-14
- [5]MassBank of North America (MoNA) · CC BY 4.0 · via PubChem · retrieved 2026-10-14
- [6]NIST Mass Spectrometry Data Center · Linked only · via PubChem · retrieved 2026-10-14
- ·Calculated values: computed by Athanor with RDKit 2026.09 from the structure.
- ·Literature on PubChem
Data last updated 2026-10-14.