
Bilirubin
Athanor · official
C33H36N4O6 · 584.7 g/mol
IUPAC name: 3-[2-[[3-(2-carboxyethyl)-5-[(4-ethenyl-3-methyl-5-oxopyrrol-2-ylidene)methyl]-4-methyl-1H-pyrrol-2-yl]methyl]-5-[(4-ethenyl-3-methyl-5-oxopyrrol-2-ylidene)methyl]-4-methyl-1H-pyrrol-3-yl]propanoic acid
Also called jaundice, bile pigment, heme breakdown
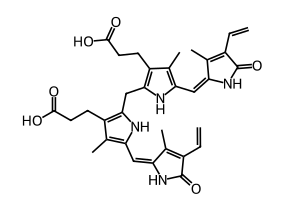
Names and identifiers
- IUPAC name
- 3-[2-[[3-(2-carboxyethyl)-5-[(4-ethenyl-3-methyl-5-oxopyrrol-2-ylidene)methyl]-4-methyl-1H-pyrrol-2-yl]methyl]-5-[(4-ethenyl-3-methyl-5-oxopyrrol-2-ylidene)methyl]-4-methyl-1H-pyrrol-3-yl]propanoic acid
- Synonyms
- jaundice, bile pigment, heme breakdown
- Molecular formula
- C33H36N4O6
- Empirical formula
- C33H36N4O6
- Molecular weight
- 584.7 g/mol
- Exact mass
- 584.26348
- CAS number
- 635-65-4
- PubChem CID
- 59687179
- Canonical SMILES
- CC1=C(NC(=C1CCC(=O)O)CC2=C(C(=C(N2)C=C3C(=C(C(=O)N3)C=C)C)C)CCC(=O)O)C=C4C(=C(C(=O)N4)C=C)C
- Isomeric SMILES
- CC1=C(NC(=C1CCC(=O)O)CC2=C(C(=C(N2)C=C3C(=C(C(=O)N3)C=C)C)C)CCC(=O)O)C=C4C(=C(C(=O)N4)C=C)C
- InChI
- InChI=1S/C33H36N4O6/c1-7-20-16(3)26(36-32(20)42)13-24-18(5)22(9-11-30(38)39)28(34-24)15-29-23(10-12-31(40)41)19(6)25(35-29)14-27-17(4)21(8-2)33(43)37-27/h7-8,13-14,34-35H,1-2,9-12,15H2,3-6H3,(H,36,42)(H,37,43)(H,38,39)(H,40,41)
- InChIKey
- OQNSGJHDCGJNHV-UHFFFAOYSA-N
- Molfile
Names, InChI and the identifiers from PubChem, retrieved 2026-10-14.
Calculated
- Heavy atoms
- 43
- Rings
- 4
- Aromatic rings
- 2
- Fraction sp³ carbons (Fsp3)
- 0.27
- H-bond donors
- 6
- H-bond acceptors
- 4 computed here · PubChem 6
- Rotatable bonds
- 12
- TPSA (Ų)
- 164.4 computed here · PubChem 164
- Formal charge
- 0
- XLogP3
- 2.9 PubChem
- Complexity
- 1240 PubChem
- Stereocenters
- None
- Double-bond geometry
- None specified
- Point group
- C1 of one 3D conformer with its hydrogens, by Athanor’s engine (as in the app)
Elemental analysis
- C67.79%
- H6.21%
- N9.58%
- O16.42%
Isotope pattern
Each peak is a nominal mass with its abundance-weighted exact mass, as a percentage of the tallest, from IUPAC natural isotope abundances.
Computed from the structure with RDKit 2026.09 when this page was built, beside PubChem's own computed values from its record; where the two differ, both are shown.
Predicted
- log P
- 4.53 Crippen (Wildman and Crippen 1999)
- log S (water solubility)
- -5.7 ESOL (Delaney 2004), log mol/L
- Synthetic accessibility
- 3.94 SA score, 1 easy to 10 hard (Ertl and Schuffenhauer 2009)
Drug-likeness: Lipinski’s rule of five
- Molecular weight ≤ 500: 584.67
- log P ≤ 5 (Crippen): 4.53
- H-bond donors ≤ 5: 6
- H-bond acceptors ≤ 10: 4
Breaks 2 of the four rules.
¹³C NMR shifts, predicted
Predictions, not measurements; each names its method. A measured log P, where a source reports one, is under Physical properties.
Safety
No GHS hazard classification was found in the sources.
Always read the supplier’s safety data sheet before working with it.
Literature
Reading PubChem, OpenAlex and GSRS…
Bioactivity
Reading ChEMBL…
References
- ·Calculated values: computed by Athanor with RDKit 2026.09 from the structure.
- ·Literature on PubChem
Data last updated 2026-10-14.